Code
Data
Paper & Results
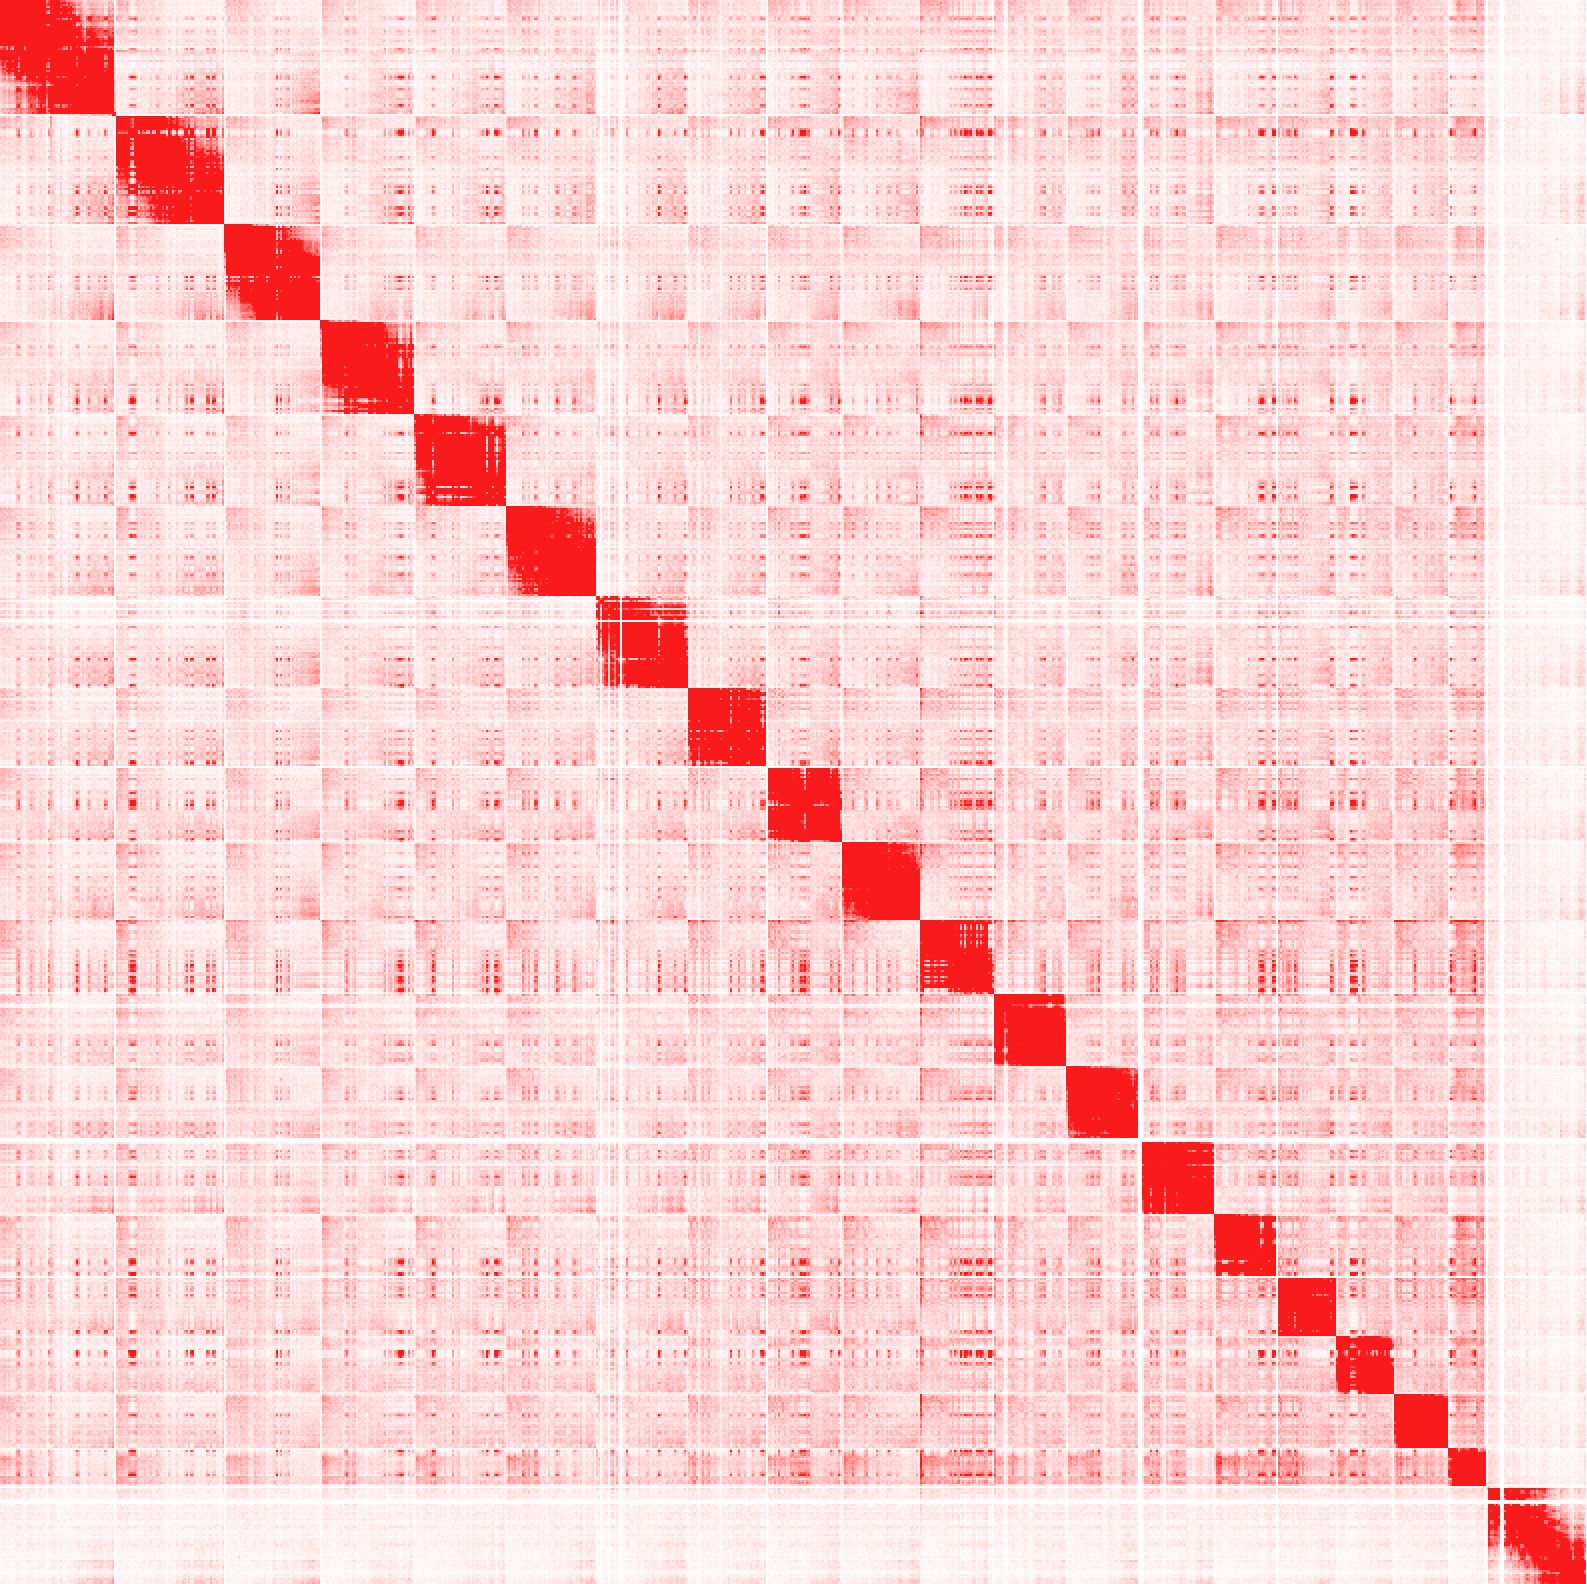
Chromosome conformation capture (3C) experiments provide a
window into the spatial packing of a genome in three
dimensions within the cell. This structure has been shown to
be highly correlated with gene regulation, cancer mutations,
and other genomic functions. However, 3C provides
measurements on a population of typically millions of cells,
each with a different genome structure due to the fluidity
of the genome and differing cell states. Thus, 3C
measurements are the convolution of measurements from
populations of cells.
We present several algorithms to
deconvolve these measured 3C matrices into estimations of
the contact matrices for each subpopulation of cells and
relative densities of each subpopulation. We formulate the
problem as that of choosing matrices and densities that
minimize the Frobenius distance between the observed 3C
matrix and the weighted sum of the estimated subpopulation
matrices. Results on HeLa 5C and mouse and bacteria Hi-C
data demonstrate the methods' effectiveness. We also show
that domain boundaries from deconvolved matrices are often
more enriched or depleted for regulatory chromatin markers
when compared to boundaries from convolved matrices.
 Download
the paper
 Reach us
Copyright © 2014 - All Rights Reserved - Emre Sefer